


OMIM ID:
Retinitis Pigmentosa 1
Alternate Names
Defective Genes
Clinical Characteristics
Ocular Features
Night blindness, the predominant presenting symptom, is often noted in the first decade of life but may not be a significant complaint until the third decade. Concentric peripheral field loss likewise follows a similar timeline. ERG responses progressively decrease in amplitude and may become undetectable in the second decade. The retinal disease progresses relentlessly, albeit slowly, as the result of photoreceptor degeneration and most patients have severe visual handicaps by midlife but there is considerable clinical variation. The pigmentary retinopathy is typical for classical retinitis pigmentosa with vascular attenuation, perivascular bone-spicule pigment clumping, optic atrophy, and generalized retinal atrophy with relative sparing of the macula early in the disease. Lens opacities are common in late stages of the disease.
Systemic Features
No systemic disease is associated with the ocular disorder caused by mutations in RP1.
Genetics
Inheritance
Multiple heterozygous, homozygous, and compound heterozygous mutations in the RP1 gene (8q12.1), sometimes called the oxygen-regulated photoreceptor protein 1 or ORP1 gene, are responsible for this disorder. The protein product is active specifically in retinal photoreceptors. Retinitis pigmentosa 1 is generally considered to be an autosomal dominant disorder and accounts for 5-7% of dominantly inherited RP disease. However, recent reports suggest that some mutations in RP1 are responsible for familial cases transmitted in an autosomal recessive pattern in which the clinical disease is more severe.
More than 20 different mutant genes have been associated with autosomal dominant RP but many cases lack a family history suggesting additional genetic heterogeneity remains. Reduced penetrance and variable expressivity characteristic of genetic disease likely contributes to the clinical heterogeneity as well. For more about autosomal dominant retinitis pigmentosa, see Retinitis Pigmentosa, AD (180380, 268000).
Pedigree
Autosomal dominant
Autosomal dominant disorders require only one mutation for the disease to be expressed. Since an affected parent has two chromosomes, only one of which has the mutant gene, parents can expect that half (50%) of their children will receive that one and inherit the disease. It is common for individuals that inherit the mutation, however, to not have evidence of the disease (nonpenetrance).
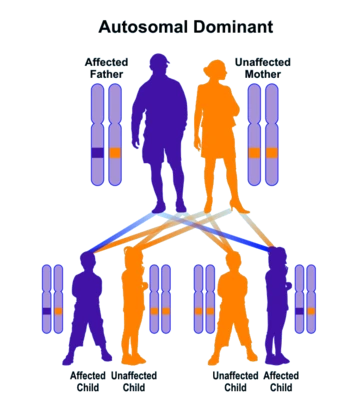
Autosomal dominant inheritance leads to a vertical pattern of transmission
Autosomal recessive
In order for autosomal recessive disorders to be expressed, offspring generally must inherit two mutations, one from each carrier parent. Carriers with only one mutation, such as the parents, do not have clinical disease. Note that carrier parents can expect that 1 in 4 children (25%) will inherit both mutations and have the disorder, 2 in 4 children (50%) will be carriers like their parents, while 1 in 4 children (25%) inherit neither mutation.

In order for autosomal recessive disorders to be expressed, offspring generally must inherit two mutations, one from each carrier parent. Carriers with only one mutation, such as the parents, do not have clinical disease. Note that carrier parents can expect that 1 in 4 children (25%) will inherit both mutations and have the disorder, 2 in 4 children (50%) will be carriers like their parents, while 1 in 4 children (25%) inherit neither mutation.